businessHosted by Aurevia & Regulatory Affairs Professionals Society (RAPS)
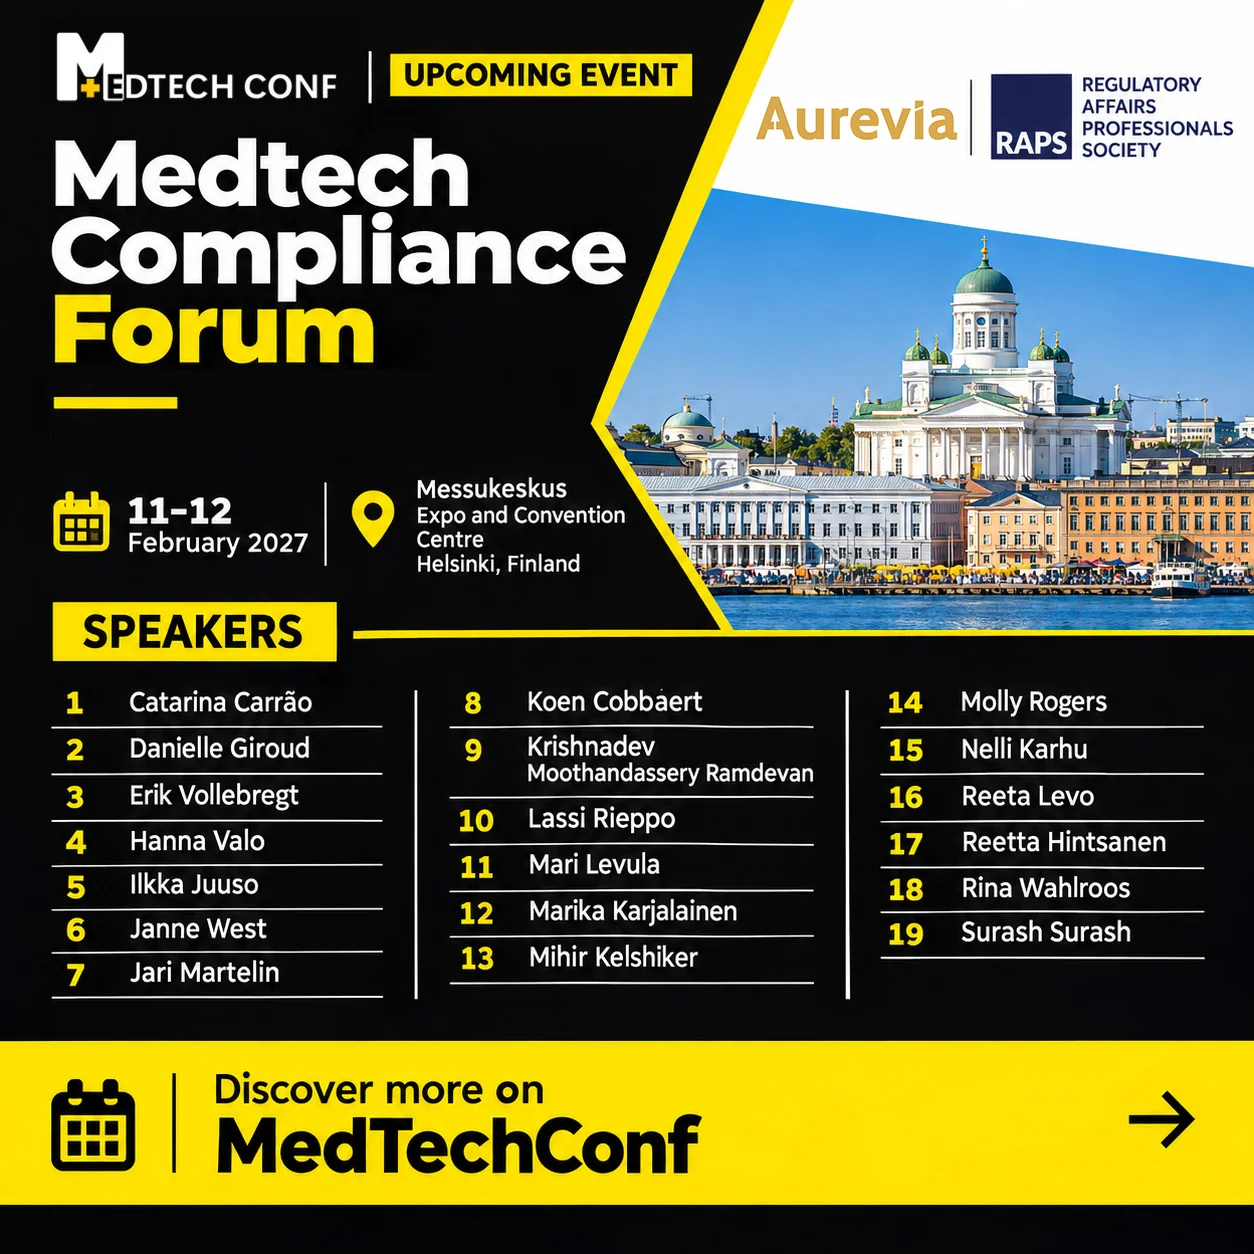
The Medtech Compliance Forum by Aurevia & RAPS takes place as part of Labquality Days 2027 in Helsinki, Finland. The conference brings together medical device and IVD manufacturers, regulatory affairs specialists, quality professionals, notified body experts and healthcare technology developers. The program explores EU MDR and IVDR, medical device software classification, AI governance, clinical evidence, clinical investigations, post-market monitoring, PMCF and PMPF, the European Health Data Space, cybersecurity documentation and regulatory market access.
Date
Time
08:00 AMVenue
Helsinki
Finland
Hosted By
Aurevia & Regulatory Affairs Professionals Society (RAPS)
Catarina Carrão
Regulatory Clinical Evaluation & Assessment, BioSciPons, Austria
Danielle Giroud
Chief Executive Officer, MD-CLINICALS, Switzerland
Erik Vollebregt
Helsinki, Finland | Messukeskus, 1, Messuaukio, Itä-Pasila, Pasila, Central major district, Helsinki, Helsinki sub-region, Uusimaa, Mainland Finland, 00520, Finland
Attorney-at-law/Partner, Axon Lawyers, The Netherlands
Hanna Valo
Fimea
Ilkka Juuso
Lead Auditor, C&T Manager, SGS Fimko (NB 0598), Finland
Janne West
Senior Quality & Regulatory Affairs Consultant, Aurevia, Sweden
Jari Martelin
Clinical Director, Bittium Medical, Finland
Koen Cobbaert
Regulatory Science & Policy Expert, Philips, Belgium
Krishnadev Moothandassery Ramdevan
Senior Quality and Regulatory Consultant, Aurevia, Sweden
Lassi Rieppo
FIMEA
Mari Levula
SGS
Marika Karjalainen
Quality & Regulatory Affairs Expert, Aurevia, Finland
Mihir Kelshiker
Clinical Scientist, Scarlet, UK
Molly Rogers
Head of IVDR, Medical Europe, DNV, Ireland
Nelli Karhu
Healthtech Finland
Reeta Levo
Bioretec
Reetta Hintsanen
Chief Operating Officer, Gosta Labs, Finland
Rina Wahlroos
VP, Quality and Regulatory Compliance, Aidian, Finland
Surash Surash
Senior Clinical Reviewer, TÜV SÜD, UK
All times are in Europe/Helsinki.
Registration and exhibition networking.
Opening of the congress.
The future of laboratory medicine through automation and AI.
MDR classification and regulatory requirements for digital health technologies.
Clinical investigation planning and execution for medical devices.
Governance and ongoing monitoring of healthcare AI.
Regulatory applications of real-world and digital clinical evidence.
Regulatory acceptance of simulated clinical evidence.
Managing AI model changes and performance throughout the medical device lifecycle.
Common clinical evaluation report deficiencies identified by notified bodies.
Clinical evidence expectations for AI-enabled medical devices.
Post-market clinical and performance evidence for medical device software and AI.
Regulatory and clinical considerations for real-world testing of medical technologies.
Clinical evaluation consultation procedure requirements.
Morning registration and exhibition.
Navigating medical technology regulatory pathways.
Explore other relevant upcoming events.

ClinChain 2027 – A Clinical Trial Supply Chain Conference is the definitive global forum for supply chain leaders and innovators dedicated to transforming the planning, execution, and manag...

The 10th edition of the Global Clinical Trials Connect Conference is a premier platform bringing together global leaders, clinical researchers, regulatory experts, and technology pioneers t...

The 10th edition of the Digi-Tech Pharma & AI 2027 Conference is a leading platform uniting pharmaceutical and technology leaders, healthcare innovators, regulatory authorities and solution...

Pharmacovigilance World 2027 is the definitive platform for global drug safety professionals, regulators, and industry innovators to connect, learn, and shape the future of pharmacovigilanc...
IVDR expectations for laboratory-developed in-house tests.
Classification of medical device software under EU MDR Rule 11.
Performance evaluation requirements for software IVDs.
Regulatory strategies for scaling medical AI products.
Applying European Health Data Space requirements to software development.
Potential implications of IVDR revisions for diagnostics manufacturers.
Software bills of materials and medical device software design.
Managing legacy IVD transition requirements.
Practical AI applications and verification in clinical workflows.
Maintaining IVDR certification and preparing for follow-up audits.
End of the congress.